





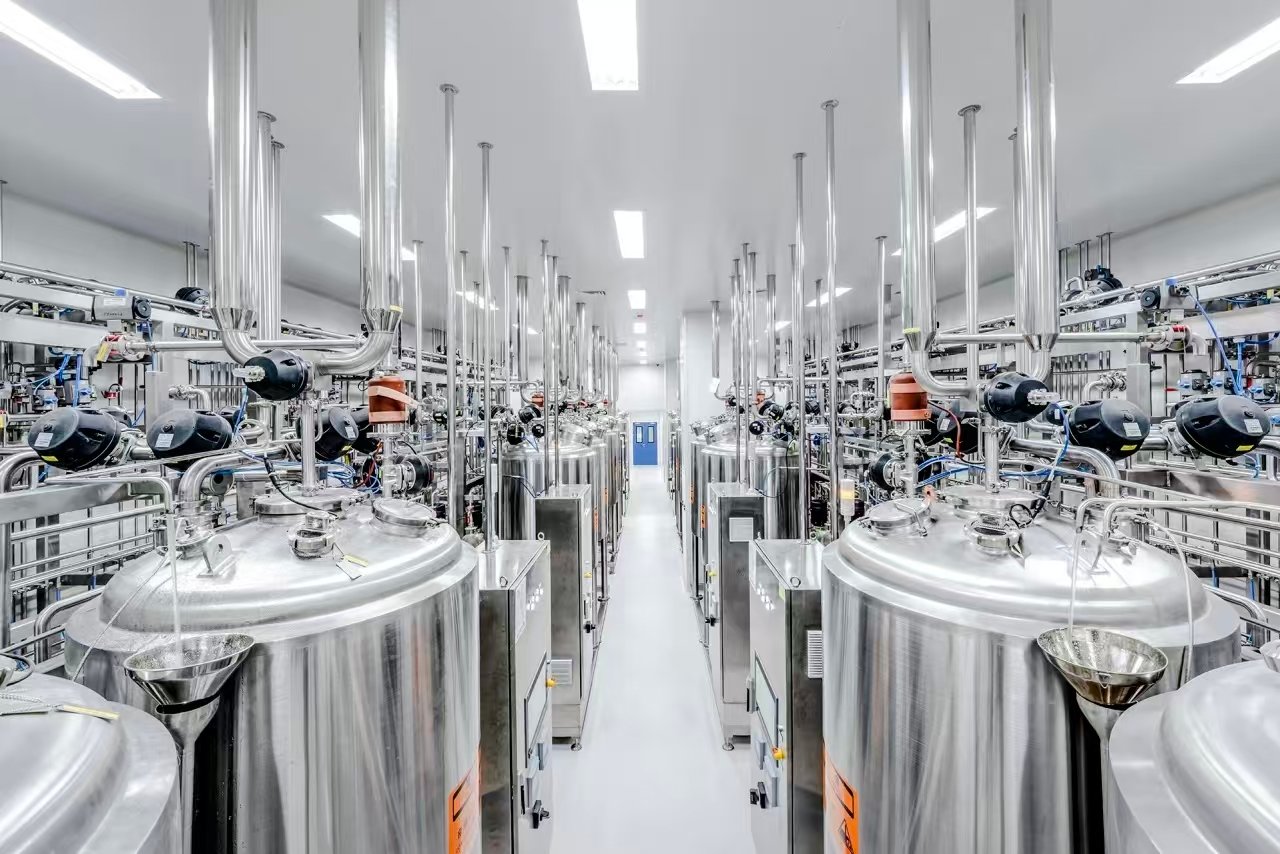
Against the backdrop of global integration in the pharmaceutical industry, overseas registration by domestic pharmaceutical manufacturers and localised production of international pharmaceutical companies have become prevailing industry trends. As a core component of the pharmaceutical quality management system, the Contamination Control Strategy (CCS) stands as a key focus during regulatory audits and product registration inspections conducted by drug authorities worldwide. The core value of the CCS system lies in systematically and structurally managing risks to comprehensively control microbial contamination, particulate contamination, cross-contamination and environmental pollutant risks throughout pharmaceutical production, so as to guarantee consistent, safe and uniform drug quality. At present, China GMP, EU GMP and FDA cGMP constitute the three primary global pharmaceutical regulatory frameworks. Distinct disparities exist between the three systems in terms of the defined framework, control dimensions, validation criteria and audit priorities for CCS. Most international pharmaceutical enterprises face prevalent challenges including applying a single CCS system to multiple regional markets, inaccurate regulatory benchmarking, blind spots in control provisions, and incomplete compliance remediation. Such issues may easily trigger regulatory inspection observations, rejection of product registration applications and production suspension risks. Based on global pharmaceutical compliance governance logic, this paper conducts an in-depth benchmarking of core CCS requirements under the three major regulatory frameworks, systematically dissects divergent points, identifies compliance loopholes, and puts forward targeted and implementable optimisation and remediation solutions. It provides technical support and practical guidance for pharmaceutical enterprises to establish a globally compliant standardised CCS system adaptable to multi-jurisdictional audits.
PART 1 Regulatory Origin and Divergent Positioning of CCS Compliance Control
Differences in the regulatory origin and positioning of CCS compliance control represent the fundamental underlying distinction among the three systems, and serve as the primary basis for enterprises to establish compliant systems. In terms of regulatory positioning, EU GMP imposes the most advanced and systematic control over CCS. Annex 1 for the Manufacture of Sterile Medicinal Products explicitly defines CCS as an independent mandatory core module of the quality management system. Enterprises are required to establish documented, end-to-end and dynamically updated CCS documentation systems covering the full lifecycle including facility design, equipment selection, production operations, environmental monitoring, cleaning and disinfection, and deviation management. CCS constitutes a prerequisite for sterile production compliance; the absence of a compliant CCS system may directly result in a non-conformity finding against the production system.
The FDA cGMP framework does not contain standalone regulatory clauses dedicated to CCS. Requirements for contamination control are scattered across multiple regulatory documents including 21 CFR Part 211, Guidance for Industry: Sterile Drug Products Produced by Aseptic Processing, Process Validation Guidance and Quality Risk Management Guidance. Its core logic is outcome-oriented, risk-driven and evidence-based. While no fixed documentation structure is mandated, all contamination control activities must be supported by complete data and risk justifications, with full implementation records retained. Greater emphasis is placed on the authenticity and effectiveness of on-site execution rather than documentary form, with extremely rigorous requirements for practical implementation.
China GMP together with the revised Annex for Sterile Medicinal Products draws extensively on the EU regulatory framework and formally incorporates CCS into mandatory control requirements. Manufacturers of sterile medicinal products and high-potency medicinal products must establish robust contamination control strategies. Compared with EU GMP’s comprehensive refined control and FDA’s outcome-oriented governance, CCS regulation under China GMP is undergoing continuous standardisation and refinement. It retains mandatory requirements for systematic documentation, while aligning with the practical status of domestic pharmaceutical production. Inspectors focus on documentation completeness, procedural standardisation and consistency between documentation and on-site practices, with steadily increasing scrutiny of dynamic risk iteration and data traceability. These fundamental disparities in regulatory positioning lead to divergent compliance priorities for enterprises in document development, on-site control, validation frequency and audit preparation.
PART 2 Disparities in CCS Documentation Structure and System Establishment
Marked differences exist between the three regulatory frameworks regarding CCS documentation architecture and system development. EU GMP specifies a unified standardised structure for CCS documents. A complete CCS system must encompass nine core modules: risk assessment reports, contamination source identification lists, detailed control measures, equipment control plans, personnel control specifications, cleaning and disinfection systems, environmental monitoring programmes, deviation and change control, and annual review reports. All modules are mandatory and must form closed-loop logic with interconnected and traceable data. Furthermore, CCS documents shall be customised according to production line type, product toxicity classification and sterility level; generic templated documents are prohibited. Separate dedicated CCS control systems are required for high-potency sterile lines, general sterile lines and solid dosage form manufacturing lines.
No fixed CCS documentation template is stipulated under the FDA framework. The primary requirements are risk-aligned, practically effective and fully traceable. Enterprises may develop CCS documentation structures independently based on production processes, product risks and facility layout without adhering to predefined modules. Nevertheless, all contamination control measures must be supported by scientific risk assessments, and all control activities shall be backed by authentic production data, monitoring data and validation data. FDA auditors focus less on whether documentation is complete or structurally standardised, and more on consistency between documented requirements and on-site operations, effectiveness of control measures in mitigating contamination risks, and integrity of raw data. “Paper compliance” where documentation diverges from on-site practices is strictly prohibited.
Documentation requirements for CCS under China GMP largely benchmark the EU architecture. Mandatory core deliverables include contamination source identification, risk grading, control measures, validation and confirmation, and continuous review. Compared with EU GMP, domestic inspections place stricter judgements on non-compliance such as non-standard documentation, missing clauses and ambiguous procedures. However, requirements for refined adaptation to differentiated scenarios and in-depth dynamic risk iteration remain relatively lenient. This constitutes a major source of inspection observations for domestic manufacturers targeting overseas markets.
PART 3 Divergences in Contamination Risk Assessment and Graded Control
Variations in contamination risk assessment and graded control represent a critical differentiator across the three frameworks, which directly determines the suitability of control measures. EU GMP mandates the deployment of standardised risk assessment tools such as FMEA and HAZOP to quantitatively grade four categories of risks: microbial contamination, particulate contamination, cross-contamination and active substance leakage. Clear control frequencies, acceptance criteria and validation requirements are defined for high, medium and low risk tiers. For high-potency and high-toxicity sterile medicinal products, separate dedicated risk assessments are required with escalated control standards, implementing the principle of “one assessment per production line, one strategy per product”, delivering highly granular risk governance.
FDA governance prioritises scientific risk judgement without mandating specific risk assessment tools. Enterprises may select appropriate assessment methodologies in light of process characteristics. However, risk assessments must be sufficiently scientifically justified through comprehensive analysis of historical deviation data, product quality data, industry risk cases and equipment operational records. The FDA attaches great importance to identification and control of rare and latent risks. Control requirements for routine low-risk scenarios are relatively flexible, while verification of control effectiveness for high-risk operations is subject to stringent scrutiny to prevent superficial, overly conservative or insufficient risk assessments.
Risk assessment under China GMP centres on standardisation and normalisation. FMEA is widely adopted with risk grading based on severity, occurrence and detectability, delivering clear and unified control logic. Nevertheless, risk assessments conducted by many domestic pharmaceutical enterprises suffer from templated and formalised practices. Risk grading poorly matches actual production scenarios, latent contamination risks are inadequately identified, and dynamic risk updates lag behind. This fails to meet the refined risk control requirements of EU GMP and FDA, representing a key area for multi-jurisdictional compliance remediation.
PART 4 Disparities in Equipment Control and Environmental Control Implementation
Noticeable gaps exist in inspection criteria for equipment governance and environmental control implementation within the CCS framework across the three regulations. EU GMP imposes stringent requirements on the containment, automation and sterilisation efficiency of cleanroom equipment, utility systems and control equipment. Mandatory deployment of fully enclosed isolator systems, automated disinfection equipment and negative pressure control equipment is required for high-risk operations, with open high-risk operations prohibited. Rigorous quantitative standards are specified for HVAC systems, pressure differentials, air change rates, VHP decontamination efficacy and filter integrity testing. Clear specifications are established for environmental monitoring frequency, sampling locations and data retention, highlighting the supporting role of hardware infrastructure in executing the CCS.
The FDA focuses on the practical contamination control performance of equipment and environments. Deployment of high-end automated equipment is not mandatory, provided that equipment consistently meets contamination control needs with complete validation data and stable operational status. Core inspection priorities include sustained equipment maintenance, stable sterilisation efficacy and authenticity of environmental data. Falsification of equipment parameters, backdating of monitoring data and superficial validation reports are prohibited. Compliance judgement of hardware equipment is solely based on actual operational performance.
Equipment control standards under China GMP benchmark EU quantitative indicators, defining acceptance criteria for cleanroom environmental parameters, equipment sterilisation standards and disinfection equipment configuration. Inspections focus on complete equipment qualifications, compliant parameters, intact maintenance records and standardised operations. Governance covers both hardware configuration and on-site execution. However, many enterprises achieve compliant hardware installation yet inadequate routine maintenance and loose parameter management, resulting in gaps against international high-level compliance expectations.
PART 5 Disparities in Validation Systems and Dynamic Review Mechanisms
Validation frameworks and dynamic review mechanisms constitute another key differentiated module of CCS compliance across the three jurisdictions. EU GMP mandates a dynamic iteration mechanism for CCS systems featuring quarterly reviews and comprehensive annual reviews. All environmental monitoring data, deviation events, change records, equipment malfunctions and product quality anomalies shall be incorporated into review scope. Risk assessments, control measures and workflows shall be optimised promptly based on review outcomes to ensure continuous alignment between the CCS system and evolving production conditions. Regular challenge testing for cleaning and disinfection efficacy, sterilisation processes and environmental control systems is compulsory with rigorous, fixed-frequency validation standards.
No fixed review frequency for CCS is specified by the FDA. The core principle is risk-triggered dynamic updates. CCS review and risk re-assessment must be initiated immediately upon production line process changes, product switching, quality deviations, equipment modifications or environmental anomalies, followed by updates to control strategies. If no abnormal risks occur, the existing system may remain in place, provided fully traceable operational data are retained to demonstrate sustained system effectiveness. The underlying logic is “no iteration without anomalies; mandatory remediation upon risk identification”, eliminating formalised periodic reviews.
China GMP explicitly requires enterprises to conduct annual CCS reviews and generate formal review reports. In practice, annual reviews performed by many enterprises feature homogenised content, incomplete data collation, unclosed corrective actions and ineffective strategy optimisation. Reviews become formalised exercises incapable of enabling dynamic optimisation of the CCS system, failing to satisfy audit requirements of EU and FDA authorities.
PART 6 Key Compliance Remediation Measures
Based on the core disparities among the three regulatory frameworks and multi-market compliance demands of pharmaceutical enterprises, targeted key compliance remediation measures are summarised to build a universally applicable standardised CCS system.
1. Upgrade of Documentation System
Enterprises shall abandon templated documentation designed solely for domestic compliance. The nine core modules shall be enhanced in line with EU GMP standardised architecture, while supplementing scientific risk justifications, on-site implementation records and full-spectrum data retention to satisfy FDA’s outcome-orientation and traceability requirements. A composite CCS documentation system shall be established featuring regulatory-aligned architecture, precise risk mapping, traceable implementation and dynamic iteration, concurrently meeting the requirements of all three regulatory jurisdictions.
2. Refined Remediation of Risk Control
Move beyond templated risk assessment practices. Customised risk assessments shall be performed for different production lines, products and processes with quantitative risk grading and targeted identification of latent contamination risks. Establish a dynamic risk update mechanism to optimise contamination source lists and control measures in real time incorporating production changes, deviation data and industry cases, balancing standardisation under EU GMP, scientific reasoning required by the FDA and normalisation stipulated by China GMP.
3. On-site Implementation and Equipment Compliance Remediation
Benchmark EU hardware control standards to address deficiencies in containment systems, automated disinfection equipment and negative pressure protection for high-risk operations. Standardise control of core parameters including HVAC systems, cleanroom pressure differentials and environmental monitoring. Comply with FDA requirements for practical execution to eliminate paper-only compliance, strengthen routine equipment maintenance, parameter management and data authenticity control, ensuring full consistency between hardware facilities, operational workflows, documented requirements and monitoring data.
4. Remediation for Iteration Mechanisms and Audit Preparedness
Implement a dual mechanism combining periodic reviews and risk-triggered iteration: conduct comprehensive annual reviews and quarterly special reviews, while enabling immediate CCS updates in response to process changes, quality deviations and equipment abnormalities. Optimise validation systems and conduct regular challenge testing for disinfection efficacy, environmental control and equipment integrity, retaining complete validation data to meet inspection standards of drug authorities across jurisdictions, eliminating barriers to multi-regional compliance.
PART 7 Conclusion
In summary, the three major regulatory frameworks including China GMP, EU GMP and FDA cGMP adopt divergent governance logic, documentation structures, risk criteria and validation requirements for CCS. EU GMP prioritises systematic standardisation and granularity; the FDA emphasises practical execution effectiveness and data integrity; China GMP focuses on procedural normalisation and documentation completeness. For pharmaceutical enterprises expanding globally, only by accurately understanding regulatory disparities, abandoning single-region compliance mindsets, and advancing system upgrading, risk refinement, on-site remediation and dynamic iteration can a globally compliant CCS system adaptable to multi-jurisdictional audits be established. This effectively mitigates compliance risks, removes barriers to market access at home and abroad, enables standardised, internationalised and high-end governance of pharmaceutical production quality, and lays a solid compliance foundation for global corporate expansion.
